


For decades, researchers have relied on gene expression to help decipher how genetic variation contributes to disease. However, what if significant genetic effects take place before changes in gene expression can be identified? And, if so, how many potential therapeutic targets have been overlooked as a result?
A new study published in Nature Genetics raises these questions by demonstrating that chromatin accessibility profiling can uncover thousands of genetic regulatory associations that conventional expression-based analyses fail to detect.
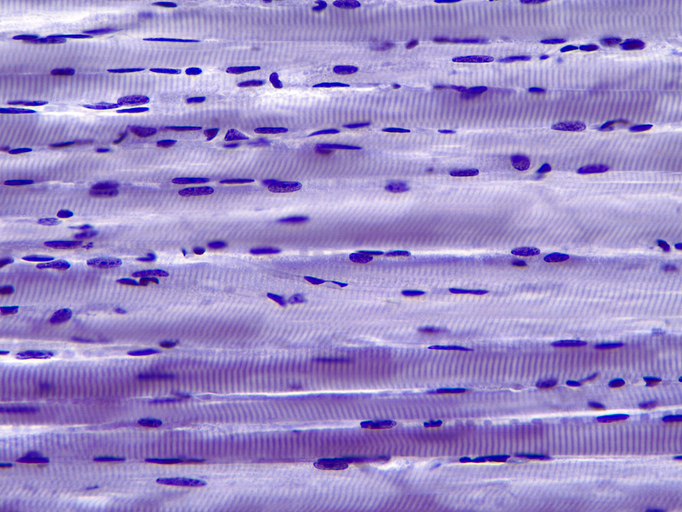
Researchers led by Arushi Varshney, PhD, and colleagues at the University of Michigan generated a population-scale, single-nucleus multiomic atlas of human skeletal muscle, integrating transcriptomic and chromatin accessibility profiles from 456,880 nuclei across 287 biopsies. The findings expose an extensive layer of cell-specific genetic regulation, offering new insights into the molecular architecture of type 2 diabetes and other complex metabolic traits.
Beyond the transcriptome
Genome-wide association studies (GWAS) have identified thousands of variants associated with metabolic disease. Yet translating these associations into biological mechanisms remains challenging because approximately 90% reside in noncoding regions of the genome.
Expression quantitative trait locus (eQTL) analyses have become a standard approach for connecting these variants to candidate effector genes. However, gene expression represents only one layer of genetic regulation. Variants may alter transcription factor binding or chromatin accessibility without producing detectable changes in steady-state RNA abundance. Such effects may become apparent only in particular cell types or under specific physiological conditions.
To investigate these regulatory layers, the researchers combined single-nucleus RNA sequencing with chromatin accessibility profiling using ATAC sequencing, resolving 13 distinct cellular populations within skeletal muscle. Integrating these measurements with donor genotypes revealed 7,062 eQTLs and 106,059 chromatin accessibility QTLs (caQTLs).
Although this approximately 15-fold difference partly reflects the greater number of chromatin features tested, the findings suggest that accessibility measurements can capture regulatory effects that remain undetectable at the transcriptomic level. Notably, caQTLs exhibited stronger genetic effects in distal enhancers, whereas eQTL effects were more prominent near transcription start sites. This distinction is particularly relevant because disease-associated genetic variants frequently map to distal regulatory elements.
Diabetes mechanisms uncovered
Integrating the atlas with GWAS datasets revealed 3,487 colocalized genetic association signals across 38 traits. Strikingly, approximately 80% colocalized exclusively with caQTLs rather than eQTLs. The implication is significant: relying solely on expression-based genetic mapping may leave many disease-associated regulatory mechanisms unresolved.
Moreover, nearly half of type 2 diabetes GWAS–caQTL colocalizations were specific to individual cell populations, highlighting the importance of cellular context. At the ARL15 locus, researchers identified a diabetes-associated regulatory signal in fibro-adipogenic progenitors, cells involved in muscle maintenance and regeneration.
Chromatin interaction analyses implicated FST, encoding follistatin, as a potential effector gene. Follistatin influences muscle growth, fat mass and insulin resistance, suggesting a possible connection between tissue remodeling and metabolic disease. Another diabetes-associated locus, C2CD4A/C2CD4B, showed regulatory activity in muscle fibers, implicating VPS13C as a candidate target gene. A massively parallel reporter assay provided experimental evidence of enhancer activity at the implicated regulatory element, although its endogenous effects remain to be established.
Rethinking genetic target discovery
The researchers used statistical causal inference to integrate chromatin accessibility and expression associations and found 1,061 molecular association pairs with consistent regulatory directionality. These findings suggest that genetic variants can affect transcription by altering chromatin accessibility, while transcription can also affect structure. Statistical colocalization and regulatory directionality do not prove molecular causality, and experimental validation of proposed mechanisms is needed.
No evidence is found that chromatin accessibility profiling directly identifies more therapeutically actionable targets than conventional methods. However, it challenges a key functional genomics assumption: that gene expression changes adequately reflect disease-associated genetic variation.
This has implications for drug discovery. Genetic associations that defy mechanistic interpretation may be functional and invisible to molecular measurements. Instead of replacing transcriptomics, population-scale chromatin profiling may help identify disease-relevant regulatory elements, cellular contexts, and candidate effector genes.
Based on these findings, it seems that the issue has shifted from determining which genes show disease-related differential expression to determining which genetic regulatory mechanisms have gone undiscovered due to the initial focus on changes in expression.